TIPS: Pharmaceutical GMP Compliance
Returned and Salvaged Drug Products Are a High-Risk Disposition Decision. These 7 Tips Keep Your Programme GMP-Compliant.
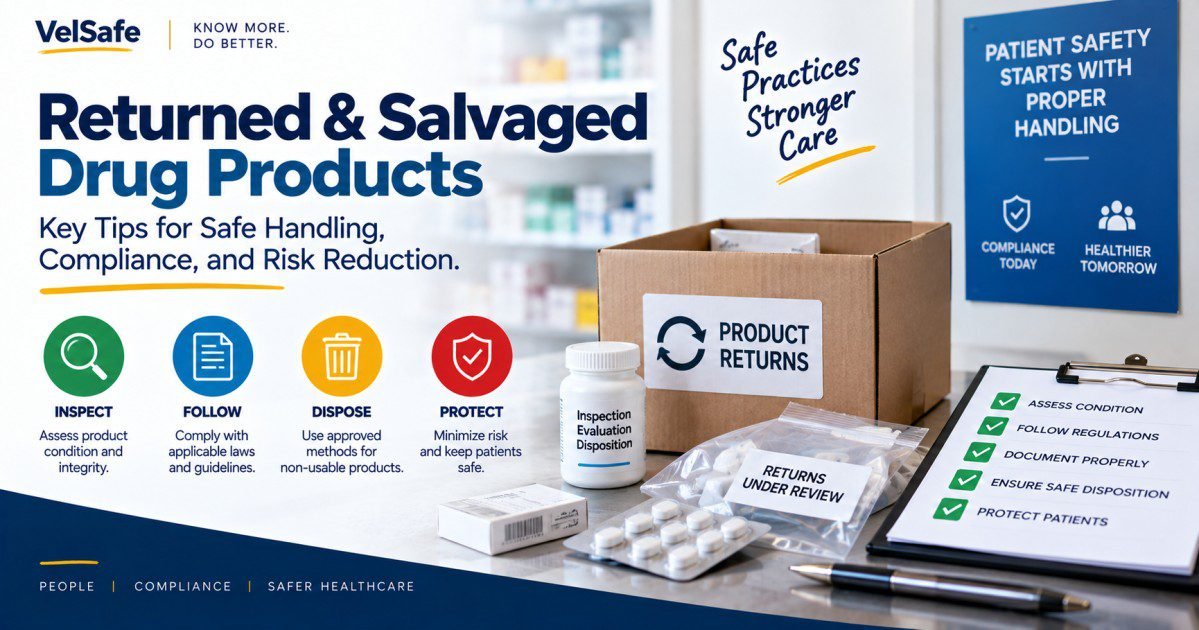
Returned and salvaged drug products process requirements under 21 CFR 211.204 are among the most frequently mishandled areas in pharmaceutical GMP programmes. FDA expects manufacturers to evaluate returned goods against documented criteria, store them under controlled conditions, and make defensible disposition decisions based on evidence, not on commercial pressure. Salvaged drug products recovered after extreme conditions (floods, fires, power failures, temperature excursions) carry additional risk and additional scrutiny. These 7 tips cover the process requirements, the documentation standards, and the GMP failures that most commonly attract FDA observations in this area.
211.204
The 21 CFR Section Governing Returned Drug Products in US GMP Regulations
21 CFR 211.204 requires that returned drug products be identified, stored under quarantine, evaluated against established criteria, and either reprocessed or destroyed based on that evaluation. The regulation prohibits returning salvaged products to the marketplace without documented evidence that quality has not been compromised. Source: 21 CFR 211.204
Quarantine
All Returned Drug Products Must Be Held Under Quarantine Until Disposition Decision Is Made
FDA requires that returned drug products be separated from released inventory and held in a designated quarantine area until evaluation is complete and a documented disposition decision is made. Returned products that enter the active inventory without quarantine and evaluation represent a serious GMP failure, and a patient safety risk if compromised product reaches the supply chain. Source: 21 CFR 211.204
Destroy
Default Disposition for Salvaged Products Is Destruction Unless Quality Can Be Demonstrated
For salvaged drug products recovered after conditions that may have compromised quality (floods, fires, power failures, extreme temperature excursions), FDA’s default expectation is destruction. Release requires documented evidence that the product has not been compromised, not merely the absence of visible damage. The burden of proof is on the manufacturer to demonstrate quality, not on FDA to prove its absence. Source: 21 CFR 211.204
Quick Summary: 7 Tips for Returned and Salvaged Drug Products
1
Quarantine all returned products immediately on receipt, before any evaluation begins
2
Document the reason for return and chain of custody before accepting the product
3
Evaluate returned products against written, pre-established disposition criteria, not case by case
4
Apply more stringent controls to salvaged products, assume compromise unless evidence proves otherwise
5
Ensure reprocessing decisions are supported by validated processes and quality unit approval
6
Maintain complete and attributable records for every returned and salvaged product disposition
7
Review returned product data periodically as a signal of potential supply chain or manufacturing quality issues
The 7 Tips: Returned and Salvaged Drug Products Process Requirements
Tip 1: Quarantine All Returned Products Immediately on Receipt
WHY IT MATTERS
21 CFR 211.204 requires that returned drug products be held in a designated area, separate from released inventory, until their disposition is determined. A returned product that bypasses quarantine and re-enters the active supply chain without evaluation is an adulteration risk. FDA inspectors frequently check returned goods procedures and verify that physical quarantine controls match written SOPs.
WHAT TO DO
Designate a physical quarantine area for returned drug products that is clearly labelled, access-controlled, and separate from released inventory. Assign status labels to all returned containers at the point of receipt. The quarantine SOP should specify who may receive returned goods, what information must be collected at receipt, and how to initiate the evaluation process within a defined timeframe.
COMMON MISTAKE
Warehouses that store returned goods in a general “hold” area shared with rejected materials, raw materials, or other quarantined inventory. Without a dedicated returned goods quarantine area and clear status labelling, there is no reliable way to distinguish returned product from other materials, and FDA will note the absence of adequate segregation.
PRO TIP
Set a maximum quarantine time in your SOP, 30 days is a common standard. Products that remain in quarantine beyond that limit without a disposition decision should trigger an escalation. Aged returns that sit in quarantine without resolution are a sign that the evaluation process has broken down.
Tip 2: Document the Reason for Return and Chain of Custody Before Accepting
WHY IT MATTERS
The reason a product was returned, who handled it after it left the manufacturer’s control, and what conditions it was stored under during the return period are all material to the disposition evaluation. A product returned because it was stored in a vehicle in extreme heat is a different risk profile from a product returned because of a labelling dispute. Without this information at the point of receipt, the evaluation will be based on incomplete data.
WHAT TO DO
Create a returned goods receipt form that captures: the product, lot number, quantity, and condition at receipt; the stated reason for return; the name and contact of the party returning the goods; the approximate date of distribution and the date of return; and any known conditions during the return transit period. This form becomes part of the returned goods record and supports the disposition evaluation. Do not accept returned goods without completing this form.
COMMON MISTAKE
Accepting returned goods from a logistics carrier or distributor without any accompanying documentation. When the evaluation team later reviews the product, they have no information about why it was returned or how it was handled post-distribution. This gap means the evaluation must either be conducted on visual inspection alone, which is inadequate, or the product must be destroyed due to insufficient information to support release.
PRO TIP
Include a section in your customer and distributor agreements specifying what documentation must accompany any returned product. A return without required documentation should be refused at the receiving dock or conditionally accepted with automatic destruction as the default disposition pending receipt of the required information.
Tip 3: Evaluate Against Written, Pre-Established Disposition Criteria
WHY IT MATTERS
21 CFR 211.204 requires that returned drug products be evaluated using established criteria. Evaluations conducted on a case-by-case basis without written standards are not GMP-compliant, they produce inconsistent decisions and are indefensible during inspection. If the disposition criteria exist only in the evaluator’s head, FDA will note the absence of a written procedure and the inability to demonstrate consistency across evaluations.
WHAT TO DO
Write a returned goods evaluation SOP that defines: the criteria for release back to stock (if permitted by product type); the criteria that require reprocessing before release; the criteria that require destruction; and the testing requirements for each disposition pathway. Criteria must be objective, based on product-specific quality attributes, and approved by the quality unit. Every disposition decision must reference the applicable written criteria and show the basis for the decision made.
COMMON MISTAKE
SOPs that list general factors to consider (“product condition”, “reason for return”, “time out of distribution”) without defining what constitutes pass or fail for each factor. Vague criteria produce inconsistent decisions, the same product could be released by one evaluator and destroyed by another. FDA’s expectation is that criteria are specific enough to produce the same decision regardless of who conducts the evaluation.
PRO TIP
Build a disposition decision tree into your SOP, a flowchart that walks the evaluator through each criterion in sequence and produces a documented decision. Decision trees reduce evaluator discretion, increase consistency, and make it easy to demonstrate during inspection that every evaluated return was assessed against the same criteria in the same order.
Tip 4: Apply Heightened Controls to Salvaged Drug Products
WHY IT MATTERS
Salvaged drug products, those recovered after exposure to conditions outside their approved storage parameters (floods, fires, power failures, extreme temperature excursions), represent a fundamentally different risk category from routine returned goods. The nature and extent of potential compromise may not be visible on inspection. FDA has specifically cited manufacturers for releasing salvaged products without adequate evidence that product quality was not affected by the compromising event.
WHAT TO DO
Establish a separate written procedure for salvaged products that specifies: the types of events that trigger salvage controls; the immediate quarantine and isolation requirements; the testing and evaluation requirements before any salvaged product may be released; and the documentation requirements for the salvage event, the product’s exposure history, and the basis for any release decision. Absent documented evidence of no quality compromise, the default disposition must be destruction. Source: 21 CFR 211.204
COMMON MISTAKE
Treating salvaged products under the same procedure as routine returned goods. The commercial pressure to recover product value after a facility event (a warehouse flood, a cold chain power failure, a fire in an adjacent area) can lead quality teams to apply the standard returned goods evaluation when the situation requires a more rigorous, event-specific assessment. FDA distinguishes between routine returns and salvage situations, and your procedures must too.
PRO TIP
For temperature-sensitive products, consider contacting the manufacturer’s stability programme team before making any salvage disposition decision. A product’s approved stability data may include excursion studies that define how long out-of-range temperatures can be tolerated, and that data can support or preclude release depending on what the exposure history shows.
Tip 5: Require Quality Unit Approval for Every Reprocessing Decision
WHY IT MATTERS
Reprocessing a returned drug product requires that there is a validated, approved reprocessing method, that the reprocessed product will meet all specifications, and that the quality unit has reviewed and approved both the decision to reprocess and the specific method to be used. Reprocessing that bypasses the quality unit is a GMP violation regardless of whether the final product meets specification. The quality unit’s approval authority over disposition decisions is a non-negotiable GMP requirement under 21 CFR 211.22.
WHAT TO DO
Document every reprocessing decision in a written record that includes: the identity and lot of the product to be reprocessed; the reason for return and the evaluation findings; the specific reprocessing method to be used and its validation status; the testing plan for the reprocessed product; and the signature of the quality unit approver. Do not initiate reprocessing operations until the quality unit approval is documented. After reprocessing, testing must confirm that all specifications are met before release. Source: 21 CFR 211.188
COMMON MISTAKE
Operations teams initiating reprocessing based on verbal quality unit approval, with the written documentation added afterward. Post-hoc documentation of quality decisions does not demonstrate that the quality unit’s approval was obtained before the operation began. FDA looks at timestamps, if the reprocessing batch record was initiated before the quality unit approval was signed, the sequence is a GMP failure regardless of the outcome.
PRO TIP
Build reprocessing into your change control or deviation management system rather than handling it as a standalone process. Linking the reprocessing decision to a deviation record or CAPA provides an audit trail that connects the quality event to the corrective action and makes the full decision history visible in a single record chain.
Tip 6: Maintain Complete, Attributable Records for Every Disposition
WHY IT MATTERS
Every returned and salvaged drug product disposition must be documented in a way that is complete, attributable, legible, contemporaneous, and original (ALCOA principles). A disposition decision made without a complete record is indefensible during inspection. FDA inspectors request returned goods records as a routine part of GMP inspections, the ability to produce a complete, accurate record for every returned lot is a basic expectation.
WHAT TO DO
Every returned goods record must include: product identity, lot number, quantity, and NDC or product code; date and source of return; reason for return as stated by the returning party; condition of the product at receipt; results of the evaluation against disposition criteria; the disposition decision (release, reprocess, or destroy) with supporting rationale; the name, title, and signature of the quality unit approver; and the date of disposition. Destruction records must include the method and witnessed confirmation of destruction.
COMMON MISTAKE
Returned goods log entries that record only the disposition decision (“destroyed”, “returned to stock”) without the evaluation findings and rationale that support the decision. A log that shows the outcome but not the basis is not a GMP record, it cannot demonstrate that an evaluation was performed, that criteria were applied, or that the quality unit approved the decision.
PRO TIP
Link your returned goods records to your batch record system so that any returned lot can be traced to its original batch record, distribution history, and complaint record if one exists. This cross-referencing is not required by regulation but makes investigations significantly faster and demonstrates a mature quality management system to FDA investigators.
Tip 7: Use Returned Product Data as a Quality Signal in Your Review Programme
WHY IT MATTERS
Returns data is a quality intelligence asset. A pattern of returns for the same product, lot, or distribution channel can signal a manufacturing defect, a packaging failure, a cold chain breakdown, or a labelling problem, before those issues escalate to a recall or a regulatory action. 21 CFR 211.180(e) requires that annual product reviews include an evaluation of returned goods data. Treating returns as isolated administrative events rather than as data in a quality surveillance programme misses the signal.
WHAT TO DO
Include returned goods data in your periodic product review (PPR/APR) process. Analyse return rates by product, lot, distribution channel, customer, and stated reason for return. Look for patterns, multiple returns for the same reason, returns concentrated in a specific geographic area, or returns from a particular distributor. Any pattern that suggests a systemic quality issue should trigger a formal investigation. Source: 21 CFR 211.180(e)
COMMON MISTAKE
Annual product reviews that include a return count (“7 returns in the calendar year”) without trending analysis or investigation of whether the returns indicate a systemic issue. A count without analysis is not an evaluation. FDA expects the PPR to demonstrate that returns data was examined for signals and that any signals identified were investigated. A rising return rate with no corresponding investigation is a red flag.
PRO TIP
Set alert thresholds in your returns tracking system, for example, any product with more than a defined number of returns in a rolling 90-day period, or any product where the return rate exceeds a defined percentage of distribution. Alert thresholds trigger a review before the annual PPR and allow you to detect and respond to emerging quality signals in real time.
Pre-Disposition Checklist: Returned and Salvaged Drug Products
At Receipt
Returned goods receipt form completed: product, lot, quantity, reason, source
Product placed in quarantine area and labelled with quarantine status
Physical condition of containers documented at receipt
Evaluation process initiated within SOP-defined timeframe
During Evaluation
Evaluation conducted against written disposition criteria in approved SOP
Salvage controls applied if event exposure is identified or suspected
Required testing performed before any release or reprocessing decision
Findings documented contemporaneously (not at end of evaluation)
At Disposition
Quality unit approval documented before disposition is executed
Rationale for disposition decision referenced in the record
Destruction witnessed and confirmed in writing
Data added to returns trending log for PPR analysis
Key Takeaways
Returned goods must be quarantined, evaluated against written criteria, and approved by the quality unit, in that order
21 CFR 211.204 requires each of these steps. The sequence matters as much as the steps themselves. A product released before evaluation is complete, or reprocessed before quality unit approval is documented, is a GMP violation regardless of the disposition outcome. Process sequence integrity is as important as the disposition decision.
Salvaged products require a separate, more rigorous procedure, not the standard returned goods SOP
Salvage situations involve products that may have been exposed to conditions that compromise quality in ways that are not visible on inspection. The default disposition is destruction. Release requires documented evidence of no quality compromise, not merely the absence of visible damage. A site that applies its standard returned goods procedure to salvage situations is operating below FDA’s expectation for this risk category.
Returns data is a quality intelligence signal, use it in your PPR programme, not just in individual disposition records
FDA’s expectation under 21 CFR 211.180(e) is that returned goods data is included in annual product reviews and analysed for patterns. A rising return rate for a product, a pattern of returns for the same reason, or returns concentrated in a specific channel are signals that warrant investigation. Using returns data only for individual disposition decisions and not as input to your product quality surveillance programme means you may be missing signals that could prevent a larger quality event.
Frequently Asked Questions
What does 21 CFR 211.204 require for returned drug products?
21 CFR 211.204 requires that returned drug products be held in a designated area under quarantine until their disposition is determined by a competent and responsible person. The regulation requires that returns be evaluated against established criteria before disposition, with only two outcomes permitted: return to the marketplace if quality has not been compromised and the product meets specifications, or destruction. The regulation also addresses salvaged drug products, specifying that products exposed to improper storage conditions shall not be salvaged and returned to the marketplace without evidence that product quality, safety, strength, purity, or identity has not been adversely affected. Source: 21 CFR 211.204
Can a returned drug product be released back to the marketplace?
Yes, under specific conditions. A returned drug product may be released back to the marketplace if: the evaluation demonstrates that the product’s identity, strength, purity, and quality have not been adversely affected; the product meets all applicable specifications; the quality unit has reviewed and approved the release decision; and the release decision is documented. Products that cannot be evaluated to this standard, including those where the storage and handling history after distribution cannot be verified, must be destroyed or reprocessed and re-evaluated before release. The commercial value of the returned product is not a factor in the disposition decision.
What is the difference between a returned drug product and a salvaged drug product?
A returned drug product is a product that has come back to the manufacturer or distributor from a customer, pharmacy, hospital, or other point in the distribution chain, typically because it was overstocked, mislabelled, near expiry, or no longer needed. A salvaged drug product is one that has been recovered from conditions that may have compromised its quality, floods, fires, extreme temperature excursions, power failures, or other events that expose the product to conditions outside its approved storage requirements. Salvaged products require more rigorous evaluation and face a higher evidentiary standard for release because the nature of the potential compromise is often non-visible.
What records must be kept for returned drug product dispositions?
Records for each returned drug product disposition must include: the product identity, lot number, quantity, and condition at receipt; the date and source of the return and the stated reason for return; the evaluation findings against disposition criteria; the disposition decision (release, reprocess, or destroy) with the documented rationale; the quality unit approver name, title, and signature; the date of disposition; and, for destruction, the method and witnessed confirmation. For reprocessed products, records must also include the validated reprocessing method used, the testing conducted on the reprocessed product, and the final release decision. All records must comply with 21 CFR Part 211 ALCOA documentation standards.
Are returned drug product data required in the annual product review?
Yes. 21 CFR 211.180(e) requires that annual product reviews (also called annual product quality reviews, or APQRs) include a review of returned drug products and complaints. The review must evaluate whether the returns data reveals any quality signal or trend, not merely count the number of returns. An annual product review that includes a return count without trend analysis or investigation of identified patterns does not satisfy FDA’s expectation. Return rate trends, patterns by distribution channel or customer, and the results of any investigations triggered by returns data should all be included. Source: 21 CFR 211.180(e)
Can a pharmaceutical company reprocess returned drug products?
Yes, but only under specific conditions. Reprocessing is permitted when: a validated reprocessing method exists and is approved; the evaluation of the returned product supports reprocessing as the appropriate disposition; the quality unit has approved the reprocessing decision before operations begin; and the reprocessed product is tested and meets all specifications before release. Reprocessing records must document the validated method used, the testing conducted, and the quality unit’s approval. A product that does not meet specifications after reprocessing must be destroyed, it cannot be reprocessed again indefinitely. Source: 21 CFR 211.204
What do FDA investigators most commonly cite in returned drug products inspections?
Based on FDA’s published Form 483 observations and warning letters, the most common findings in returned drug product inspections include: failure to adequately quarantine returned products separate from released inventory; disposition decisions made without written evaluation criteria; lack of quality unit approval documented before disposition; incomplete or missing returned goods records; inadequate controls for salvaged products including applying the same evaluation standard as routine returns; and failure to include returned goods data in annual product reviews with trend analysis. Sites that have these systems documented and operating consistently are well-positioned to defend their returned goods programme during inspection.
Sources
Government and Regulatory Sources
- 21 CFR 211.204, Returned Drug Products: primary regulatory source for all requirements in this article, quarantine, evaluation, disposition criteria, quality unit approval, salvaged product controls, and the prohibition on returning compromised products to the marketplace.
- 21 CFR 211.180(e), Records and Reports: Annual Product Review: source for the requirement to include returned drug products data with trend analysis in the annual product review.
- 21 CFR 211.188, Batch Production and Control Records: source for documentation requirements applicable to reprocessed returned drug products, including the requirement for quality unit approval before reprocessing begins.
- 21 CFR 211.22, Responsibilities of Quality Control Unit: source for the quality unit’s non-delegable approval authority over returned goods disposition decisions.
Related VelSafe Articles
Guides
MedTech Europe Code of Ethical Business Practice: A Guide
The European medical technology industry conduct standard covering HCP interactions, product promotion, and transparency, the marketing and commercial conduct framework that complements the manufacturing GMP requirements in this article.
Law
Medicare Part D Medication Therapy Management Law
The regulatory framework governing medication therapy management in the Medicare Part D context, relevant for pharmaceutical organisations whose returned and salvaged products include drugs dispensed under Part D plans.
Guides
Medicare Part D PDP Enrollment: A Complete Guide
The enrollment and plan operations guide for Part D, the distribution channel context for pharmaceutical companies whose returned product programmes include Part D dispensed medications and who need to understand the regulatory environment their products operate in after distribution.
PHARMACEUTICAL GMP COMPLIANCE
More Pharmaceutical GMP Tips and Guides
Explore VelSafe’s pharmaceutical GMP compliance library for tips, guides, and regulatory analysis covering 21 CFR Part 211 requirements, quality system obligations, documentation standards, and the GMP compliance areas most frequently cited in FDA inspections.
Explore All Tips